
How to choose composition analyzers for low-concentration measurements


Select composition analyzers against the concentration at which a measurement must support an action. A detection limit below that concentration is a useful starting point, but it does not establish that the instrument can quantify the substance accurately in a real sample. The decision also depends on the sample matrix, interfering compounds, sampling losses, calibration and the time available to produce a result.
Begin by defining the target substance, its chemical form and the required reporting basis. A trace amount of a dissolved metal, for example, presents a different measurement problem from the same element suspended in particles. State the concentration range, the decision threshold, the acceptable uncertainty and whether results are needed continuously or from discrete samples. These requirements give supplier specifications a common basis for comparison.
Use the quantification limit at the decision point
The limit of detection describes a level at which a signal can be distinguished from a blank according to a specified statistical procedure. The limit of quantification, or LOQ, is intended to identify a level at which measurement meets defined performance requirements. Neither is an intrinsic constant that applies to every sample and operating setup.
A detection limit derived from repeated measurements of a clean laboratory blank can look impressive while saying little about a wet process gas or a colored wastewater sample. The usable LOQ also depends on sample preparation, dilution, blank contamination, integration time and the criteria used to establish acceptable precision and bias. Ask whether the stated value refers to the instrument signal, the prepared solution or the original sample.
Place the required decision threshold above the demonstrated LOQ for the complete method, with enough room for the uncertainty associated with the decision. A result reported close to the LOQ deserves particular attention: if its uncertainty overlaps a specification limit, the measurement alone may not establish which side of the limit the true value lies on. Agree in advance on the decision rule, including how uncertainty and repeat measurements will be treated.
Units and reference conditions also matter. A concentration expressed on a dry gas basis cannot be compared directly with one reported on a wet basis. For liquids, confirm whether the result refers to mass per volume, mass fraction or another basis, and account for any dilution or extraction factor. A low instrument-level LOQ loses its advantage if sample preparation raises the reporting limit above the required threshold.
Match the measurement principle to the interference
Composition analyzers differ in what they distinguish. The choice depends on the target and the substances accompanying it, rather than on sensitivity alone. The following comparisons describe typical trade-offs; actual performance must be demonstrated for the proposed method and matrix.
A separation step is valuable when a detector responds to several compounds present in the sample. It also adds method-development work and can lengthen the interval between results. A direct optical measurement avoids chromatographic separation, but it needs a spectral region and correction strategy that remain adequate as the background composition changes. Neither approach is inherently more selective without considering the full measurement method.
For elemental analysis, identify whether the requirement concerns total concentration, dissolved concentration or a particular chemical species. An instrument that reports an element after digestion does not, by itself, establish which species was originally present. Conversely, a selective measurement of one species does not establish the total amount. Filtration, preservation, digestion or separation can therefore be part of the measurement definition, not merely preparation details.
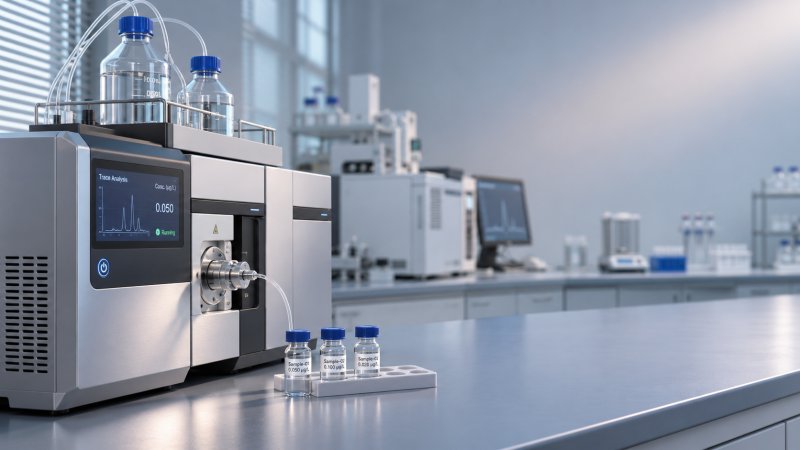
Treat the sample path as part of the analyzer
At low concentration, the sample reaching the detector may differ from the sample at the process or collection point. Adsorption onto tubing, reaction with wetted surfaces, condensation, leaks and contamination from fittings can all change the result. The importance of each mechanism follows from the target chemistry and the sample conditions.
For a gas measurement, trace moisture or a reactive contaminant can adhere to sample lines or be removed by a conditioning system. A chilled sample line can create condensate that dissolves a water-soluble analyte. A dryer that protects the sensor can also remove some of the target. Specify line materials, temperature control, pressure reduction, filtration and conditioning together, then test recovery through the complete path. A calibration gas introduced directly at the analyzer inlet verifies less of the system than one introduced near the sampling point.
Liquid samples present different difficulties. Suspended solids can settle or clog a filter; trace constituents can bind to containers; volatile compounds can be lost during transfer. If the requirement is dissolved concentration, filtration conditions belong in the method. If total concentration is required, removing particles without accounting for their contribution changes the measurand. Blank containers and preparation reagents are especially relevant when their contribution approaches the reporting limit.
Residence time also changes the usefulness of an online result. Long tubing, conditioning volume and analyzer cycle time delay the indication of a process change. Averaging reduces random noise but adds further delay and can flatten short peaks. Compare the total response at the sample point with the duration of the event that must be detected, including transport time and any filtering applied to the output.
Evaluate matrix effects with representative material
A calibration prepared in a clean solvent or balance gas does not necessarily predict response in a real sample. Salts, organic compounds, humidity, acidity and background gases can alter recovery or detector response. A stable reading is therefore not proof of an accurate reading: a consistent interference can produce a precise but biased result.
Request testing across the expected matrix range, including credible extremes rather than only a convenient average sample. Where suitable, add a known amount of target to representative material and assess the change in measured concentration. Spike recovery can reveal suppression, losses or interference, although it does not necessarily reproduce the behavior of analyte that has been bound or aged in the original sample. Where available, compare results with a suitable reference material or an independently validated method.
Investigate unexpected low results before accepting them as evidence of low concentration. A failed recovery, a rising blank, carryover from a preceding high sample and a shifted calibration response point to different causes. Sample dilution can reduce matrix interference, but it also raises the effective reporting limit for the original sample. The dilution is useful only if the remaining sensitivity still supports the decision.
Compare calibration and stability under the intended duty
A specification measured immediately after calibration does not describe behavior over an entire operating interval. Ask for evidence of zero and span stability, the conditions of the test, and the interval over which the stated drift applies. Temperature changes, optical contamination, sensor consumption and changes in sample composition can affect the result between calibrations.
Calibration standards must represent the target and concentration range being measured. For trace gas analysis, the balance gas and humidity can matter; for liquids, the solvent, acidity and matrix can influence response. A single-point adjustment may correct an offset or gain under defined conditions, but it does not establish linearity across a range. Use independent verification material where feasible, and distinguish a calibration standard from a quality-control sample used to evaluate the calibration.
Carryover deserves a separate test when high and low concentrations occur in sequence. After a high sample, residual analyte in the injection system, cell or tubing can appear as a positive signal in the next low sample. Alternating representative high samples with blanks or low-level samples exposes this behavior more clearly than repeated measurements at one concentration. The required wash or purge time must be included in the stated throughput.
For continuous service, establish what happens during calibration, cleaning and fault conditions. A system that produces a valid result only part of the time may be unsuitable for detecting short excursions, even when its laboratory sensitivity is excellent. Determine how invalid readings are flagged and whether the control system receives a current measurement, a held value or a fault signal.
Make the comparison on a common evidence package
Ask for performance data that identify the analyte, matrix, concentration, sample preparation, operating conditions and calculation method. A specification without those conditions is difficult to compare with another instrument’s claim. The most useful evidence includes replicate low-level measurements, blank results, recovery or bias data, interference studies and stability over the proposed calibration interval.
A practical acceptance trial can use representative samples and known additions at concentrations near the decision threshold. Include blanks, expected background variation and a high-to-low sequence if carryover is plausible. For an online installation, route the challenge through the proposed sampling and conditioning equipment wherever feasible. Record both result quality and the time taken to obtain a valid result after a change.
Finally, examine the uncertainty budget for the complete measurement: calibration, repeatability, bias or recovery, sample handling and any relevant drift. Avoid adding uncertainty terms mechanically when they are correlated or already included in an empirical estimate. If the demonstrated uncertainty is too large for the required decision, improving the sampling system or method may be more effective than choosing an analyzer with a lower advertised detection limit.
Recommended for You
 How to choose composition analyzers for low-concentration measurementsChoose composition analyzers with confidence. Discover how quantification limits, sample handling and matrix effects shape reliable low-concentration measurements.
 How does online analysis compare with laboratory testing for water quality monitoring?Online analysis vs. laboratory testing: compare speed, accuracy, costs, and compliance to find the right water quality monitoring strategy.
 How to choose the right detection range for a custom gas analyzerChoose the right custom gas analyzer detection range with practical guidance on accuracy, exposure limits, calibration, and supplier evaluation.
 How does multi component monitoring detect gases in stack emissions?Discover how multi component monitoring detects stack gases, manages interference and supports reliable emissions reporting. Explore key CEMS selection factors.
 How bright should an industrial monitor be for outdoor use?Choosing an outdoor industrial monitor? Learn why 1,000 nits is a starting point and how glare, contrast, and heat affect sunlight readability.
 How does water vapor affect infrared gas analyzers?Infrared gas analyzers can be affected by humidity, condensation and spectral interference. Learn how to protect sample integrity and achieve reliable results.
 How to reduce customer support response timesReduce Customer Support response times with actionable tips for reporting equipment faults, choosing the right channel, and getting expert help faster.
 How to choose a low range analyzer for trace-level measurementsChoose the right low range analyzer for trace-level measurements. Compare detection limits, sampling, calibration and support to make a confident decision.
 How to choose combustible gas detectors for a specific gasChoose combustible gas detectors with confidence. Compare sensor technologies, gas response, calibration, placement, and certification for your application.
 How to choose an air quality analyzer for the pollutants you need to measureChoose an air quality analyzer with confidence. Compare pollutant measurement methods, sampling, accuracy and calibration to find the right fit for your needs.
 How to choose industrial metrology tools for tight tolerancesChoose industrial metrology tools for tight tolerances with confidence. Compare uncertainty, calibration, tool types, and costs to reduce inspection risk.
 How moisture affects fixed analyzer accuracy in stack gas monitoringImprove fixed analyzer accuracy in stack gas monitoring. Discover how moisture causes dilution, interference, and sample loss—and learn practical ways to protect emissions data.
 How does production line speed affect in-line measurement accuracy?Discover how production line speed affects in-line measurement accuracy. Explore sensor response, sampling, and timing to boost throughput without compromising quality.
 How condensation affects HCl concentration analyzer accuracyHCl concentration analyzer accuracy can suffer when condensation traps HCl. Learn how to spot sampling losses, prevent low readings, and improve measurement reliability.
 How does cross-interference affect multi-component gas analyzers?How does cross-interference affect multi component gas analyzers? Explore measurement errors, compensation limits, and validation tips to choose the right system for your process.
 How does temperature affect high-accuracy sensor performance?Explore how temperature affects high accuracy sensor performance. Learn to reduce thermal drift, evaluate compensation, and select the right sensor for reliable measurements.
 Can a C4H8O concentration analyzer distinguish between isomers?Can a C4H8O concentration analyzer distinguish isomers? Compare GC, FTIR and MS methods, uncover selectivity limits, and learn what to validate before choosing your instrument.
 How to identify a reliable supplier using on-time delivery recordsIdentify a Reliable Supplier by verifying on-time delivery records, original commitments, and complete shipments. Learn practical checks to reduce purchasing risk and protect your project schedule.
 How sensitive should a portable hydrogen analyzer be for lab use?Choose a portable hydrogen analyzer for laboratory use with the right quantification limit, accuracy, and range. Learn how to verify performance and avoid costly buying mistakes.
 How to set industrial calibration intervals based on equipment riskSet industrial calibration intervals based on equipment risk, drift, and as-found data. Learn when to shorten or extend schedules to improve reliability and avoid unnecessary costs.
 How to choose electrochemical analyzers for low-current measurementsCompare electrochemical analyzers for precise low-current measurements. Learn how noise, leakage, bandwidth, and real-cell testing help you choose the right instrument.
 What tolerance does a custom gas mixture need for calibration?Choose the right custom gas tolerance for calibration. Learn how blending tolerance, certified uncertainty, and stability affect accuracy—and avoid paying for precision you don’t need.
 How condensation affects stack detector readings in wet flue gasStack detector readings can drift when wet flue gas condenses. Learn how to identify moisture interference, prevent sample losses, and improve emissions monitoring reliability.
 How to prevent condensation in a CO2 analyzer sampling systemPrevent condensation in your CO2 analyzer sampling system. Learn how to control dew point, eliminate cold spots, and manage moisture for reliable readings and fewer costly disruptions.
 How to verify accuracy claims for industrial quality analyzersVerify accuracy claims for industrial quality analyzers with calibration checks, independent testing, and clear acceptance criteria. Learn how to reduce risk and buy with confidence.
 How to calculate cooling loads for an industrial analyzer shelterSize industrial analyzer shelter cooling with confidence. Learn to calculate equipment heat, solar gains, and ventilation loads, then select HVAC capacity for reliable analyzer performance.
 Hydrogen Analyzers for Energy Storage: Purity Monitoring vs. Leak DetectionChoosing a hydrogen analyzer for energy storage systems? Compare purity monitoring and leak detection to select the right sensors, improve safety, and protect fuel cell performance.
 NH3 Monitoring in Fertilizer Plants: Where Should Leak Analyzers Be Installed?NH3 analyzer for fertilizer production: discover where to place leak detectors around storage, compressors, and loading areas to reduce blind spots and improve early warning.
 Diesel Engine NOx Analyzers: Response Time and NO2 Conversion ExplainedChoosing a NOX analyzer for diesel engines? Learn how response time, NO2 conversion, and sampling design affect accuracy—and make a smarter, evidence-based purchase.
 SO2 Analyzers for Smelter Emissions: Sampling Hot, Dust-Laden Flue GasSelect an SO2 analyzer for smelting industry applications. Discover how to handle hot, dust-laden flue gas, prevent sample losses, and achieve reliable emissions monitoring.
 Unstable CH4 Readings in Biogas Monitoring? Check Moisture and Sample FlowCH4 analyzer for biogas monitoring giving unstable readings? Discover how moisture, sample flow, and pressure checks help pinpoint faults and restore reliable methane measurements.
 Buying a CO2 Analyzer for Brewery Gas Recovery: Costs Beyond the Purchase PriceBuying a CO2 analyzer for brewery applications? Compare sampling, installation, calibration, and downtime costs to uncover hidden expenses and make a smarter gas recovery investment.




Search Categories
Search Categories
- Smart Meters & Power Quality
- Transformer Gas Analyzers
- BESS Thermal Runaway Detectors
- Energy, Power & Smart Grid Monitoring
- Optical Profilers & CMM
- Ultrasonic & X-Ray NDT
- Calibration Standards
- Precision Metrology & NDT
- Water Quality Analyzers
- CEMS & VOCs Monitors
- Price Trends
- Import & Export Updates
- Market Trends
- Industry News
- Industry Insights
Latest Article
- How to choose composition analyzers for low-concentration measurementsChoose composition analyzers with confidence. Discover how quantification limits, sample handling and matrix effects shape reliable low-concentration measurements.Posted by:
- How does online analysis compare with laboratory testing for water quality monitoring?Online analysis vs. laboratory testing: compare speed, accuracy, costs, and compliance to find the right water quality monitoring strategy.Posted by:
- How to choose the right detection range for a custom gas analyzerChoose the right custom gas analyzer detection range with practical guidance on accuracy, exposure limits, calibration, and supplier evaluation.Posted by:
Please give us a message
